This refers to the observation that transmission of PrPSc between two different species is typically far less efficient than transmission within the same species. This barrier may be partially due to changes in amino acid sequence, but can also be due to changes in the self-propagating structure of the protein itself. Indeed, even within a single species, pathological D-Pantothenic acid sodium variation in TSEs and different biochemical signatures of PrPSc have been observed, leading to the isolation of distinct PrPSc types. These different types of PrPSc are called prion strains, and represent amyloid conformations of PrP that are structurally unique. In many cases, different prion strains show differences in biochemical properties, such as protease resistance or denaturant sensitivity, which correlate with variation in pathology and the time course of disease. However, in other cases, prion strains have been isolated that vary in pathology, yet remain biochemically indistinguishable, according to the levels of sensitivity available with current assays. Moreover, while genetic polymorphisms in PrP bias the formation of particular conformations of PrPSc, a single primary sequence can propagate a multitude of distinct prion strains. Indeed, it has been estimated that the range of heterogeneity seen in samples from patients with sporadic CJD represents over 30 distinct prion strains. Clearly, the structural limits of amyloid polymorphism of prion strains are quite large. Interestingly, functionally distinct prion Gambogic-acid proteins exist in fungi such as the yeast Saccharomyces cerevisiae. Yeast prion proteins share many of the same misfolding and aggregation characteristics as the proteins associated with human protein conformational disorders. As such, yeast has provided a tractable model system to investigate many facets of protein aggregation and prion biology, including that of prion strain diversity. As in mammals, prion strains in yeast are conformationally distinct, self-propagating amyloid structures. This formation of amyloid in yeast leads to changes in cellular phenotypes, which typically resemble a loss-of-function phenotype of the prion protein. One of the most well-studied prion proteins in S. cerevisiae is the translation termination factor Sup35. Sup35 is the eRF3 that normally exists in a complex that functions to recognize stop codons in mRNA and facilitate the release of polypeptide chains from ribosomes. Conversion of Sup35 into its prion form,, establishes a loss-of-function phenotype that is dominant and inherited in a non-Mendelian fashion. In cells, much 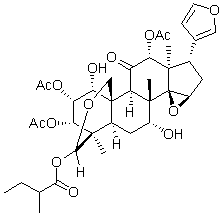 of the Sup35 is sequestered into prion aggregates, thereby impairing translation termination and causing readthrough of stop codons. variants have been broadly classified into categories based on the degree of nonsense suppression. Two wellcharacterized variants are strong and weak. Cells propagating the strong variant exhibit a greater amount of nonsense suppression as compared to cells propagating the weak variant. Studies of strong and weak led to a model that proposed an explanation for how differences in the biochemical properties of these variants correlate with differences in biological phenotypes. This model posits that decreased fiber stability results in increased fragmentation, thereby giving rise to a greater number of prion seeds, and thus more fibril “free ends” that can recruit and sequester natively-folded Sup35. Ultimately, the more ��free ends�� available are hypothesized to correlate to an increased rate of fiber growth that, in the case of the prion, modulates the strength of the nonsense suppression phenotype as the efficiency of translation termination is linked to the size of the soluble, active pool of Sup35.
Category: Kinase Inhibitors
Treatment was unable to increase the expression be beneficial in acute settings of cardiac stress
Conditions with moderate pathology, but not chronic or severe settings. Given the current enthusiasm and anticipation regarding therapeutic development of miR-34a and miR-34 family-targeted antimiRs, and the differences in cardiac protection in acute versus chronic settings, it is important to assess the therapeutic potential of inhibiting miR34a in more sustained pathological settings. This is also important because a challenge facing miRNA-based therapies is the vast number of predicted targets which could result in off-target Tubeimoside-I effects. Since the miR-34 family has approximately 31�C55% more targets in humans than miR-34a alone, interventions that modulate the entire miRNA family have greater theoretical potential to generate off-target effects. Thus, if manipulation of a single miRNA can provide adequate cardiac protection, this approach may be an advisable alternative means of obtaining a therapeutic effect with lower risk of off-target complications. In the current study, we assessed the therapeutic potential of inhibiting miR-34a in a mouse model with pre-existing pathological hypertrophy and systolic dysfunction due to pressure overload induced by TAC. We report here, that administration of a locked nucleic acid –Tulathromycin B antimiR-34a in a model of pressure overloadinduced hypertrophic cardiomyopathy with moderate systolic dysfunction was able to prevent further deterioration in cardiac function over an eight week period. In contrast, this approach was unable to attenuate pathological remodeling in a model of pressure overload with severe systolic dysfunction, whereby systolic function continued to decline eight weeks after treatment. In a previous study, we reported that inhibition of the miR-34 family attenuated LV remodeling and atrial enlargement in mouse models with established cardiac dysfunction due to MI or pressure overload. Interestingly, inhibition of miR-34a alone provided no significant benefit in the MI setting, but was not explored in the pressure overload model. Given that inhibition of miR-34a is being considered as a possible therapy for cardiovascular disease, we set out to determine whether miR-34a inhibition could provide benefit in a mouse model of pressure overloadinduced pathological hypertrophy. A potential advantage of inhibiting miR-34a alone, as opposed to the entire family, is related to fewer possible off-target effects. The major finding of the present study is that inhibition of miR34a provided some protection in the TAC moderate model, but not in the TAC severe model. LNA-antimiR-34a treated TAC moderate mice did not develop lung congestion, atrial enlargement was attenuated, and importantly, there was no further deterioration in cardiac function at 12 weeks post TAC compared to pretreatment values at four weeks post TAC. In contrast, systolic function continued to fall in TAC moderate LNA-control mice. HW/TL increased by approximately 49% in TAC moderate LNA-control mice, and only 30% in TAC moderate LNA-antimiR-34a mice. Despite attenuation of adverse cardiac remodeling, treatment with LNA-antimiR-34a was not 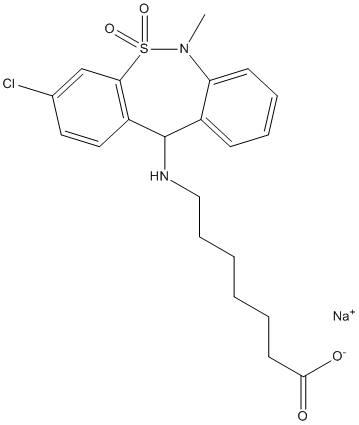 associated with a more favorable cardiac molecular signature or less fibrosis. The only parameter which showed a tendency to be attenuated in antimiR-34a treated mice was b-MHC expression. Treatment with LNA-antimiR-34a was associated with a trend for increased Sirt1 expression. Sirt1 protects the heart against ageing and stress, thus increased Sirt1 mRNA may contribute to the modest cardiac protection observed in TAC moderate mice.
Internationally comparable data with respect with two or more concurrent medical conditions are scarce
During HHcy, increased reactive oxygen species, matrix metalloproteinase, and decreased endothelial nitric oxide cause vascular constriction and stiffness, which could lead to essential hypertension. A clinical study came to the same conclusion, that Hcy was a determinant of vascular thickness and compliance in hypertensive patients and that HHcy increased the risk of cerebrovascular complications and glomerular sclerosis. There is a pathogenic effect of Hcy on the vasculature, but the data on the relationship between Hcy levels and vascular biology at the initial stages of hypertension was limited. The results of our study suggest that at an initial stage of hypertension, lower Hcy levels might indicate higher risk of the disease. The Hcy-related vascular lesion should be further investigated. A gender difference was also noted in the present study. Females had lower Hcy levels compared with males, and Hcy was not a significant risk factor for females. This difference might be due to female hormones, which were demonstrated to have Monoammoniumglycyrrhizinate antioxidant effects that may antagonize the risk based on Hcy. Further studies are needed to investigate the mechanism, but gender is an important factor that should be considered in future studies on the relationship between Hcy and hypertension. This study has some limitations. Firstly, the 2-year follow-up period might be insufficient, and could lead to bias in underestimating the influence of Hcy on the disease. However, as mentioned above, a similar approximately U-shaped curve for risk was found in a 17.5-year follow-up study, and more research or meta-analyses are needed to assess any potential bias. From 2010 to the present, we measured only two blood biochemical parameters. Although the follow-up time was only two years, we also came to the conclusion that the effect of HCY on the incidence of hypertension. The fifth physical examination will occur in 2014, and we will continue to focus on the impact of Hcy on the incidence of hypertension. Secondly, serum folate and vitamin B, which are known to influence Hcy levels, were not evaluated in this study. Thirdly, serum Hcy levels in some of the 2012�C2013 physical examinations were missing. Fourthly, we have not completed an analysis of genetic susceptibility for hypertension in our study population. In this longitudinal study, we found an approximately U-shaped risk distribution of Hcy levels for incident hypertension, especially in males. Other than the increasing risk derived from crosssectional studies and the non significant association found later in several prospective studies, we hypothesize that HHcy might  not be Danshensu Simply related to the increasing risk of incident hypertension; moreover, mildly elevated Hcy levels might be a protective factor in males. From the point of view of this study, caution should be used when considering Hcy levels to guide dietary supplementation or therapy aimed at decreasing the risk of hypertension. Simply lowering Hcy levels might not decrease the risk, or even lead to a reverse effect in some specific populations. However, we should also be aware that besides lowering Hcy levels, dietary supplementation with folate or vitamin B, both of which are in the Hcy metabolism pathway, provide additional benefits beyond the risk of incident hypertension. Rather than deny these benefits, the results of this study remind us of the possibility that there are other mechanisms that may explain how Hcy may be involved in vascular pathology.
Preserving coordinate regulation of their gene expression patterns across rice and barley in each
They encoded proteins with a wide range of biological functions. Those cBRs highly accumulated in dry mature grains and their accumulation gradually decreased over the course of seed germination. This raises the possibility that these cBRs encoded proteins involved in seed development and maturation. The highly accumulated transcripts were degraded over the course of seed germination. Barley and rice diverged from their common ancestor 50 million years ago. However, they share a great similarity morphologically and physiologically in germination and seedling growth. In this study, we measured the  transcriptomes of germinating rice grains at dry, mid- and end points of seed germinations, which should represent the most distinct stages of the dynamic transcriptional changes over seed germination process. Having determined transcriptomes of rice at the three equivalent stages, we designed a systems and evolutionary strategy to compare the dynamic transcriptomic changes over the course of seed germination to gain an insight into divergence and conservation of gene regulatory programs underlying rice and barley germination. One-Way ANOVA analysis of the transcriptomes revealed that 2537 barley and 13813 rice genes were differentially regulated over the course of seed germination. Comparing their encoding protein sequences and expression patterns identified 322 sets of conserved barley and rice genes sharing strong similarity in both protein sequences and gene expression patterns. The collection of cBRs contained 368 barley genes and 388 rice genes. Thus, only a very small percentage of the germination-regulated genes preserved their protein sequences and gene expression patterns; and a significant divergence occurred in transcriptional regulatory programs underlying rice and barley germination since the barley-rice divergence. As expected, protein sequence similarity of germination regulated barley and rice genes positively correlated to the similarity of their expression patterns, suggesting co-evolution of protein functions and gene expression patterns. Biological functions of genes are mainly determined by their protein sequences and their expression patterns. Both protein sequences and expression patterns change quickly if the genes have no Diacerein functional significance. Therefore, we hypothesized that the germination regulated expression patterns and protein sequences of the barley and rice genes in each cBR have been preserved for 50 million years after the split of rice and barley from their common ancestor because the genes are functionally important to seed germination, and should contribute to the characteristics shared by rice and barley germination. Additionally, 60 of the 322 cBRs were multi-gene cBRs. Each multi-gene cBRs contained at least one pair of paralogs. Duplicated paralogous genes are subjected to little functional constrains, and offer a great opportunity for their sub-functionalization or neofunctionalization through divergence of their protein sequence and/or expression patterns. Preserving germination regulated expression patterns and protein sequences of those paralogous genes in the multi-gene cBRs suggests that they may be subjected to negative selection, and provides additional Atropine sulfate evidence supporting their functional significance in seed germination. We identified a number of biological pathways enriched with cBRs of similar expression patterns, suggesting that their underlying transcriptional regulatory programs are highly conserved in rice and barley.
Dependent inhibitory effect of calcium on absorption of iron has been reported for molar ratios ranging
Although more recently a higher threshold molar ratio of 220:1 was reported. Our study demonstrates an inhibitory effect of calcium at a molar ratio of 11:1. There is one other human study in which the effect of alginate on iron absorption was measured. Six ileostomy patients were given a low-fiber diet with or without a daily supplement of 7.5 g sodium alginate. In 5 of the 6 subjects alginate resulted in a lower apparent absorption of iron, although the results were not statistically significant due to high inter-individual variation and insufficient power. However, as the diets were similar apart from the addition of sodium alginate, the results suggest an inhibitory effect of alginate. Bosscher et al. reported that the availability of iron and zinc from infant formulas increased with the addition of alginic acid when investigated using an in vitro dialysis model with a preliminary intraluminal digestive phase. The nature of the binding of alginate to iron is unknown. Results from circular dichroism and zeta potential experiments indicate that the interaction at pH 3.5 is through a site-binding model, with an estimated 66 sites per molecule of sodium alginate. Increasingly, amino acid changes in HA, resulting from either natural evolution or experimental design, are compared to amino acids within another subtype. A common example are those mutations that have been shown to confer binding to human glycans. In Clofentezine strains from the H3 subtype, these are Gln226Leu and Gly228Ser whereas in strains from the H5 subtype these mutations are positions 222 and 224. Although simple ‘rules-of-thumb’ can be derived, such as the subtracting four from the H3 numbering to get the position in H5 viruses, this is not  always straightforward, as typified by the recent focus on H7 viruses. The HA of H7 strains contain many amino acid insertions and deletions relative to viruses from the other subtypes. For amino acids close to the receptor binding site, such as the aforementioned mutations, the H7 numbering differs from H3 numbering by nine residues. However, two other mutations of concern, His103Tyr and Thr315Ile, which were recently shown to facilitate the aerosol transmission of avian A/H5N1 viruses between mammals, lie in the N and C termini of HA1, respectively. Due to the indels in these regions, the equivalent amino acids in H7 strains differ by three and six amino acids, respectively. As shown for H7, the conversion of residue numbering between subtypes varies depending on the region of HA being compared. Yet another complication arises due to genetic changes within a subtype which, although uncommon, do occur. Over one-fifth of the avian H5N1 strains in the Middle East sequenced to date have a deletion between amino acids positions 128 and 130. This deletion was also found in human seasonal H1 strains after 1995 but was not present in early H1 strains or any of the H1pdm strains currently circulating. Similarly, a clade of H7 strains circulating in North Ergosterol America and Canada since 1996 has been shown to have eight amino acids deleted, located surprisingly close to the receptor binding site. Conversion rules thus also depend upon the lineage of the subtypes that are being compared. Nobusawa and colleagues previously predicted the N-terminal sequence for thirteen subtypes of HA based on the likely signal peptide cleavage site of the N-terminal signal peptide, thus providing a numbering scheme based on the mature sequence of HA. Although widely cited, not all publications use this numbering. For example, only two out of the thirteen currently available crystal structures of HA of the vaccine strain of H1pdm start with the mature HA sequence. Alternative structures include six or ten additional N-terminal amino acids.